Cell Therapy Companies: Manufacturing May Become Your Biggest Bottleneck
 Written byNeta Kela, PhDFounder & Managing Partner
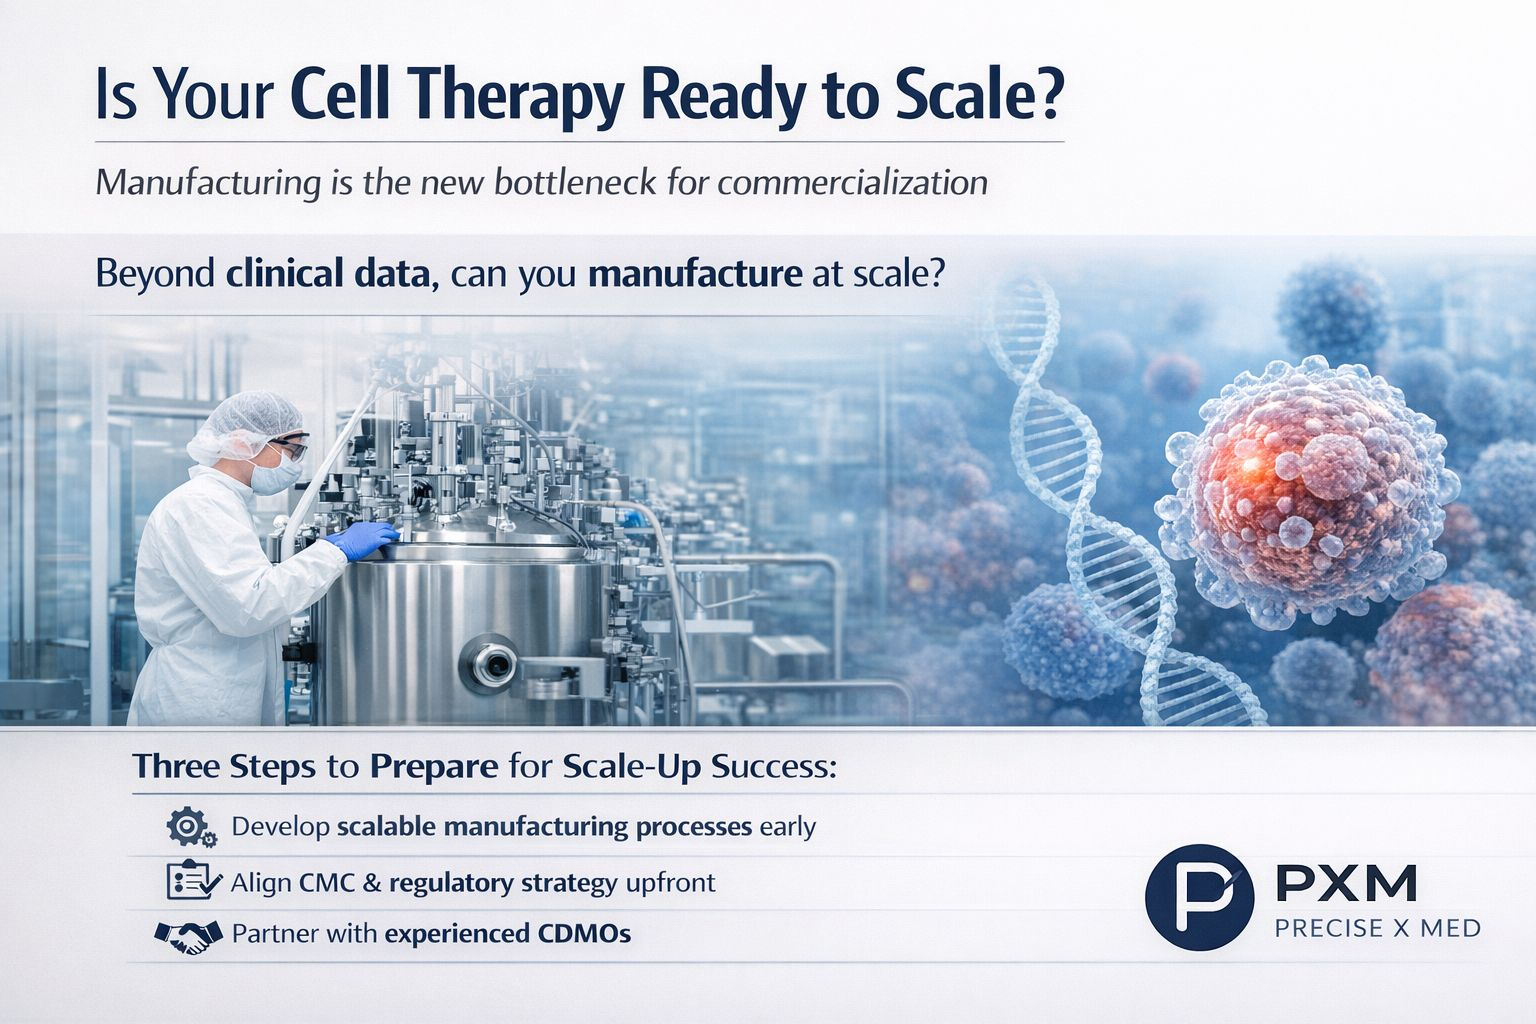
Many biotech teams focus heavily on discovery and clinical data. But regulators are increasingly asking a different question:
Can you manufacture your therapy consistently and at scale?
A recent approval from the U.S. Food and Drug Administration for a new advanced therapy manufacturing facility highlights how critical manufacturing readiness has become for commercialization.
For cell and gene therapy companies, the challenge is no longer only scientific — it is industrializing the science.
Three things biotech teams should consider early:
1. Design manufacturing with scale in mind.
Process development and facility strategy should start long before pivotal trials.
2. Align CMC and regulatory strategy early.
Manufacturing validation can become a major gating factor for approvals.
3. Choose manufacturing partners strategically.
The right CDMO or manufacturing setup can significantly reduce development risk.
Many promising therapies face delays not because of the biology — but because manufacturing was not designed early enough.
How PXM Helps With This
CMC Strategy & Manufacturing Readiness
CMC roadmaps, CDMO selection, comparability strategy, and CMC submissions.
IND-Enabling Regulatory Strategy
Pre-IND strategy, nonclinical package design, and IND readiness planning.
Biotech Readiness Assessment
A 10 minute diagnostic of your regulatory, CMC, and partnering readiness.
Related
Part of FDA & CMC Readiness. Start from the pillar: Operate practice, FDA Regulatory Readiness, CMC & Manufacturing, FDA Expedited Pathways, EU Regulatory Pathways
What a Clinical Overview Must Contain for an EU Hybrid Application
What the Clinical Overview (Module 2.5) must cover in an EU hybrid application under Article 10(3), how to justify differences from the reference product, and where assessors push back.
Well-Established Use Applications: Building a Bibliographic Clinical Overview That Holds Up
How to build the Clinical Overview for an EU well-established use application under Article 10a: the ten-year test, literature selection, and how to bridge your product to the published evidence.
How to Choose an Independent Clinical Expert for Your EU Dossier
What Article 12 of Directive 2001/83/EC requires from the Clinical and Nonclinical Expert, what to look for when you appoint one, and the questions to ask before they sign.
Get notified when we publish new funding and regulatory guides
Practical guidance on FDA and EMA strategy, CMC readiness, and non-dilutive funding — sent only when there is something worth reading.